An experimental one-time gene therapy from biotech company uniQure continues to show signs of slowing Huntington’s disease four years after treatment, offering potentially important evidence that gene therapy could alter the course of a devastating inherited neurological disorder for which no approved disease-modifying treatment currently exists.
The treatment, called ifezuntirgene inilparvovec, or AMT-130, is designed to reduce production of the huntingtin protein responsible for Huntington’s disease.
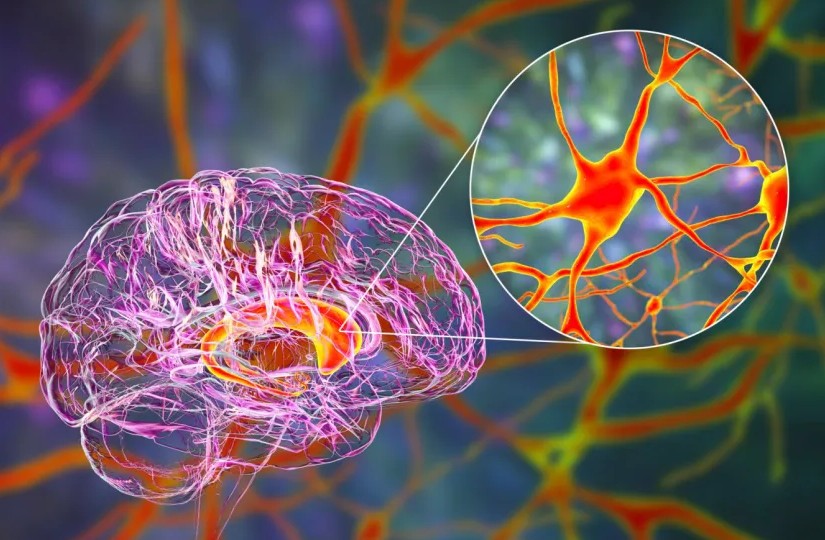
Huntington’s is caused by a mutation in the huntingtin gene and progressively damages nerve cells in the brain. Patients experience worsening movement problems, cognitive decline and psychiatric symptoms, with the disease eventually causing severe disability and death.
AMT-130 uses an engineered AAV5 viral vector to deliver genetic material directly into areas of the brain affected by Huntington’s. The therapy is administered once through stereotactic brain surgery and is intended to suppress production of huntingtin protein over the long term.
The latest analysis followed 12 patients who received the high dose for four years.
Using the composite Unified Huntington’s Disease Rating Scale, or cUHDRS, researchers calculated that disease progression was 44% slower among treated patients compared with a matched external control group drawn from a large natural-history database.
However, that difference was not statistically significant, an important limitation when interpreting the results.
Another measure, Total Functional Capacity, produced a stronger result. It indicated a 61% slowing of functional decline at four years, with a nominal p-value of 0.008.
The results nevertheless raised questions about durability because the apparent benefit was smaller than earlier findings.
At three years, uniQure had previously reported that high-dose AMT-130 slowed progression on the cUHDRS by approximately 75%. An updated analysis incorporating 15 high-dose patients now estimates an 80% slowing at 36 months.
The difference between the three- and four-year findings does not necessarily establish that the therapy is losing effectiveness.
UniQure argues that the four-year comparison was affected by missing data and changes in the external control population. A post-hoc analysis using the previous external control suggested a 54% slowing on cUHDRS at four years.
Still, the apparent reduction in treatment effect has attracted scrutiny because Huntington’s is a lifelong progressive disease. For a one-time therapy to meaningfully transform treatment, researchers will want evidence that its benefits remain substantial over many years.
The trial also has important methodological limitations.
AMT-130 has been tested in relatively small groups, and its efficacy is being compared primarily against external natural-history controls rather than a large randomized placebo-controlled trial. Such comparisons can be valuable in rare diseases but may introduce differences between treated patients and the people used as controls.
Despite those uncertainties, the findings remain notable because Huntington’s currently has treatments that can help manage certain symptoms but none approved to slow the underlying disease itself.
UniQure has already submitted applications seeking regulatory approval for AMT-130 in both the United States and United Kingdom.
The therapy’s ultimate significance will therefore depend on how regulators interpret the total evidence — including its safety, the strength of the three-year results, the less pronounced four-year findings and the limitations of comparing patients with external controls.
If future data confirm a durable effect, AMT-130 could represent an important shift in Huntington’s treatment: from managing the symptoms of an inexorably progressive neurological disease toward directly targeting its genetic cause with a single treatment designed to slow deterioration for years.